Emerging and re-emerging viruses remain among the most difficult public-health threats to anticipate, not because individual molecular events are poorly understood, but because epidemic potential emerges from interacting molecular, ecological and social processes that rarely align in predictable ways. A new narrative review published in Frontiers in Virology by Abdul Bari Hejran, Parwiz Niazi and Mohammad Yaqoub Sangin synthesises evidence published between 2017 and July 2026 across major virus families, including coronaviruses, orthomyxoviruses, filoviruses, arenaviruses, nairoviruses, hantaviruses, phenuiviruses, henipaviruses, flaviviruses, alphaviruses and orthopoxviruses. Rather than treating sequence features as deterministic predictors of emergence, the authors distinguish mechanistic plausibility from phenotypic and epidemiological validation, arguing that a systems-virology approach can improve prioritisation when predictions are explicitly graded, experimentally validated and embedded within equitable One Health surveillance.
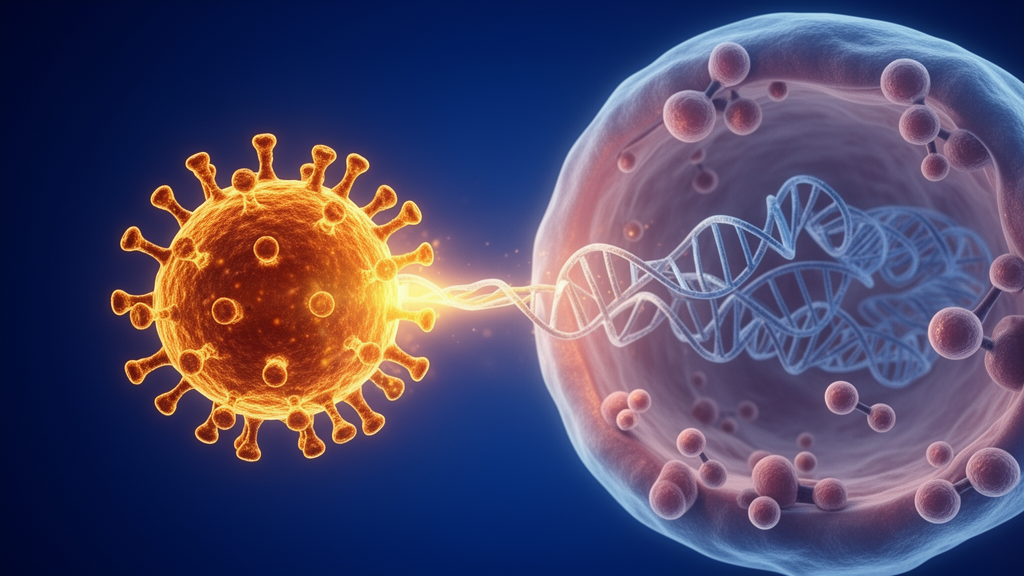
The review frames viral emergence as a cascade of linked molecular gates rather than a single event. Viral surface proteins determine attachment and entry, but compatibility between viral glycoproteins and host receptors does not guarantee productive infection. Intracellular host factors, polymerase activity, codon usage, replication organelles, interferon sensitivity and immune evasion proteins collectively determine whether progeny virus can be produced and transmitted. For SARS-CoV-2, the convergence of ACE2 engagement, spike priming by TMPRSS2, furin-site biology, replication-complex efficiency and immune evasion created a biologically favourable foundation for pandemic expansion. The same logic explains why a virus may be detected frequently at the animal-human interface yet fail to establish sustained human transmission, whereas another virus with modest ecological exposure may spread explosively once critical within-host barriers are crossed.
At the family level, the review identifies distinct molecular architectures that shape emergence risk. Coronaviruses combine large RNA genomes, recombination, broad mammalian reservoirs and spike-mediated receptor flexibility. Orthomyxoviruses such as influenza A possess segmented genomes that permit reassortment, enabling rapid antigenic and host-range shifts. Filoviruses and paramyxoviruses have non-segmented negative-sense RNA genomes, but their glycoproteins and immune-antagonist proteins facilitate severe disease in susceptible hosts. Flaviviruses and alphaviruses are shaped by vector ecology, climate-sensitive transmission and envelope-protein adaptation. Orthopoxviruses demonstrate that large DNA viruses can still undergo epidemiologically meaningful evolution under transmission pressure, as mpox has shown under new contact networks. This family-level view is not merely taxonomic; it provides a practical map for prototype-pathogen research, in which studying one coronavirus polymerase, protease or spike architecture can inform assay development and therapeutic design across related coronaviruses.
Entry is the first molecular gate through which an emerging virus must pass. Enveloped viruses use surface glycoproteins to bind receptors or attachment factors and then undergo fusion at the plasma membrane or within endosomal compartments. For henipaviruses, ephrin-B2 and ephrin-B3 receptor usage helps explain endothelial and neuronal tropism. For filoviruses, macropinocytosis, glycoprotein processing and NPC1-dependent endosomal entry are central determinants of permissiveness. However, tropism is also determined by host proteases, attachment factors, glycan presentation and innate-state context. A receptor may be expressed in a tissue yet fail to support efficient infection if the proteolytic environment or replication cofactors are absent. Conversely, viruses can exploit multiple entry routes or evolve altered receptor-binding surfaces that expand their host range. This explains why entry studies should be integrated with organoid models, primary cells, pseudovirus assays and animal-interface surveillance rather than being interpreted from receptor-binding data alone.
Replication machinery constitutes a second determinant of emergence. RNA-dependent RNA polymerases, viral proteases, nucleocapsid proteins and replication organelles determine not only viral yield but also mutation rate, recombination potential and sensitivity to nucleoside analogues. Coronaviruses are unusual among RNA viruses because their replication complex includes proofreading activity, enabling large genomes while still permitting adaptive diversity. This balance of fidelity and flexibility is one reason SARS-CoV-2 could accumulate meaningful mutations without losing genome integrity. Assembly mechanisms differ across viral families but often expose vulnerabilities: coronavirus assembly occurs at ER-Golgi intermediate compartments through coordinated interactions among M, E, S and N proteins; influenza assembly requires packaging of eight genome segments, creating opportunities for reassortment but also imposing strict compatibility constraints; filovirus budding depends heavily on matrix protein VP40 and host ESCRT machinery. Host-factor screens and proteomics have expanded the field from single viral proteins to systems-level replication networks. A SARS-CoV-2 protein-interaction map identified host pathways that could be repurposing targets, while CRISPR screens revealed TMEM41B and other host factors as pan-coronavirus dependencies.
Innate immunity imposes one of the strongest barriers to emergence, and successful emerging viruses frequently encode proteins that mask pathogen-associated molecular patterns, block adaptor proteins, degrade signalling molecules, suppress STAT activation, remodel stress responses or antagonise antigen presentation. These mechanisms are not incidental virulence accessories; they are often prerequisites for efficient replication in a new host. Convergent immune-evasion logic can be seen across unrelated viruses: influenza A NS1 inhibits antiviral RNA sensing and interferon responses; Ebola virus VP35 suppresses RIG-I-like receptor signalling while VP24 interferes with interferon-induced nuclear import; coronaviruses deploy several antagonists, including nsp1-mediated host translation suppression and accessory proteins that disrupt interferon signalling; henipavirus V, W and C proteins impair interferon production or response; and orthopoxviruses encode multiple soluble and intracellular immunomodulators. Pathogenesis results from both direct viral injury and dysregulated host response, with severe disease reflecting tropism for endothelial, respiratory, neurological or immune cells alongside excessive inflammation, vascular leakage, coagulopathy or immunopathology.
Evolutionary dynamics determine whether control measures remain effective over time. SARS-CoV-2 demonstrated that variants can accumulate spike mutations that alter transmissibility, antigenicity and neutralisation while maintaining receptor engagement. Omicron showed that immune escape can become a dominant population-level advantage in settings of widespread infection- or vaccine-derived immunity, yet escape must remain compatible with entry and replication fitness. Influenza viruses exemplify the risk of segmented genomes: reassortment between avian, swine and human viruses can generate new constellations of internal genes and surface antigens, while point mutations in HA, PB2 and other segments influence receptor preference and mammalian adaptation. Highly pathogenic H5N1 clade 2.3.4.4b has increased concern because infection in mammals creates additional opportunities for adaptation, even though sustained human-to-human transmission has not been established. The review argues that genomic surveillance should evaluate variants as phenotypic hypotheses rather than as lists of mutations, because a mutation in a receptor-binding domain, polymerase, protease, matrix protein or immune antagonist may be important only in a specific genomic background.
Rapid diagnostics are central to containment, but performance is conditional. RT-qPCR is sensitive and scalable, yet primer mismatch, sampling time, specimen quality and infrastructure can reduce effectiveness. Programmable CRISPR assays can shorten redesign cycles, whereas metagenomic next-generation sequencing offers agnostic detection when the causative agent is unknown, divergent or coinfecting; neither replaces prospective clinical validation, contamination control or external quality assurance. Outbreak genomics has transformed reconstruction for Ebola, Zika, yellow fever, SARS-CoV-2 and mpox by linking sequence data to transmission dynamics, revealing introductions, hidden transmission, superspreading and international dissemination. The value of genomic data depends on representative sampling, metadata quality, rapid deposition, analytic capacity and equitable participation by affected regions. On the countermeasure side, the COVID-19 pandemic demonstrated both the power and fragility of platform approaches: mRNA, adenoviral-vector and protein-based vaccines can be developed quickly when antigen design, manufacturing capacity, clinical trial networks and regulatory pathways are prepared in advance, but antigenic evolution can reduce neutralisation, requiring booster strategies, updated antigens or broader immunogen design. Direct-acting antivirals are most useful when they target conserved enzymatic functions, have acceptable safety profiles and can be delivered early in infection, as remdesivir and nirmatrelvir-ritonavir showed in selected clinical settings, while molnupiravir demonstrated more modest clinical utility and raises different resistance and mutagenesis considerations.
For filoviruses, the PALM trial established that monoclonal antibody therapies can improve outcomes for Ebola virus disease when delivered with supportive care, shifting the field from empiricism towards evidence-based outbreak therapeutics. For mpox, tecovirimat has been widely used because of orthopoxvirus rationale and animal-rule approval, but recent human trial evidence for clade I mpox did not demonstrate faster lesion resolution, highlighting the need for outbreak-embedded randomised trials rather than reliance on plausible mechanism alone. The evidence base remains structurally uneven: SARS-CoV-2 offers abundant structural, clinical and population data, while Lassa virus, Crimean-Congo haemorrhagic fever virus, hantaviruses and Rift Valley fever virus remain comparatively under-characterised. This asymmetry can make publication volume appear equivalent to biological importance. Lassa-virus diversity and immune evasion complicate vaccine generalisation; CCHFV combines tick ecology with incomplete correlates of protection; hantavirus pathogenesis centres on endothelial dysfunction despite limited specific therapy; and RVFV risk spans livestock, vectors and human disease. Conflicting evidence commonly arises from model choice, because receptor overexpression can exaggerate permissiveness, pseudoviruses isolate entry while omitting replication and immune antagonism, organoids improve tissue relevance but incompletely reproduce systemic immunity, and animal models may not capture human transmission.
The review proposes a deliberately modular sequence-to-decision standard in which a novel signal passes through five stages: detection and metadata review; family and structural triage; functional testing of entry, replication and innate responses; explicit evidence grading; and a proportionate decision about surveillance, assay updating, countermeasure screening or clinical research. Artificial intelligence can accelerate protein-structure prediction, variant prioritisation, epitope screening and surveillance anomaly detection, yet confidence scores do not establish biological activity, host range or clinical relevance, and training-data bias, poor representation of disordered complexes and limited prospective validation require that AI outputs remain auditable hypotheses tested against experimental and epidemiological data. Across virus families, three implementation priorities recur: assay standardisation so that variant phenotyping does not depend on isolated laboratories using incomparable methods; predefined connections between molecular evidence and public-health thresholds, such as repeated detection of a high-risk virus in a new host or convergent mutations at receptor-binding sites; and sustainable capacity in genomic sequencing, biobanking, bioinformatics, clinical trials and vaccine delivery that must exist before a crisis, especially in regions where spillover and neglected outbreaks are most likely to occur. Molecular systems analysis cannot predict emergence with certainty, but evidence-graded integration can shorten the interval between a credible signal and a proportionate response while reducing both false reassurance and unjustified alarm.
Subject of Research: Molecular mechanisms of viral emergence, immune evasion and cross-species adaptation across major virus families, with a sequence-to-decision framework for precision countermeasures
Article Title: Molecular systems of emerging viruses: replication, host immune evasion, cross-species adaptation and precision countermeasures
Article References: Molecular systems of emerging viruses: replication, host immune evasion, cross-species adaptation and precision countermeasures. (n.d.). https://doi.org/10.3389/fviro.2026.1926007
Image Credits: AI Generated
DOI: 10.3389/fviro.2026.1926007
Keywords: emerging viruses, systems virology, SARS-CoV-2, immune evasion, cross-species transmission, genomic surveillance, CRISPR diagnostics, vaccine platforms, antivirals, One Health, zoonotic spillover, viral replication
Cite Scienmag News
Kristina Jarvis. (October 16, 2026). How Emerging Viruses Cross Species Barriers: A Molecular Systems View of Replication, Immune Evasion and Precision Countermeasures. Scienmag. https://scienmag.com/how-emerging-viruses-cross-species-barriers-a-molecular-systems-view-of-replication-immune-evasion-and-precision-countermeasures/
Kristina Jarvis. "How Emerging Viruses Cross Species Barriers: A Molecular Systems View of Replication, Immune Evasion and Precision Countermeasures." Scienmag, 16 October 2026, https://scienmag.com/how-emerging-viruses-cross-species-barriers-a-molecular-systems-view-of-replication-immune-evasion-and-precision-countermeasures/. Accessed 16 October 2026.
Kristina Jarvis. "How Emerging Viruses Cross Species Barriers: A Molecular Systems View of Replication, Immune Evasion and Precision Countermeasures." Scienmag. October 16, 2026. https://scienmag.com/how-emerging-viruses-cross-species-barriers-a-molecular-systems-view-of-replication-immune-evasion-and-precision-countermeasures/