

Dr. Michael F. Peintinger from the Mulliken Center for Theoretical Chemistry at the University of Bonn has developed vibe-qc, a general-purpose quantum chemistry program designed to compute the electronic structures of molecules and crystals, using generative artificial intelligence agents to write and test the codebase. Published in the Journal of Computational Chemistry, the project demonstrates how an automated programming system guided by a scientific researcher can construct a functional computational chemistry framework from the ground up in approximately ten weeks. Rather than introducing a novel individual calculation formula, the work establishes a development methodology in which autonomous digital assistants draft code modules and execute unit tests under direct scientific oversight.
The foundational motivation for quantum chemical software centers on the calculation of electronic structures, as the spatial distribution and behavior of electrons dictate virtually all macroscopic and microscopic properties of matter. These electron arrangements govern whether a bulk solid conducts electricity, the specific wavelengths absorbed to give a dye its color, the mechanical stability of crystalline lattices, and the kinetic rates at which chemical transformations proceed. Because electrons do not follow rigid deterministic trajectories, quantum chemistry applies the principles of quantum mechanics to determine spatial probability distributions that define where electrons are most likely to reside across atomic nuclei.
To illustrate practical chemical modeling, Peintinger examined the surface adsorption of water molecules onto a wafer-thin layer of boron nitride, a compound synthesized from alternating boron and nitrogen atoms. In such a computational simulation, researchers evaluate how strongly a molecule adheres to a substrate and determine its preferred geometric position by placing the adsorbate at multiple coordinates, varying its spatial orientation, and calculating the relative total energies of the resulting configurations. The structural arrangement yielding the lowest overall energy represents the thermodynamically most favorable state, a determination that provides critical physical insight into the mechanisms governing surface-mediated catalytic reactions.
Despite the clarity of quantum mechanical principles, fundamental physical constraints complicate their mathematical execution in multi-electron systems. Analytical solutions to the governing equations can be obtained exactly only for single-electron configurations, such as an isolated hydrogen atom, whereas even a microscopic crystal of diamond contains countless mutually interacting electrons whose correlated behavior cannot be solved without systematic approximations. Computational chemists must therefore select approximation techniques tailored to the specific accuracy requirements and computational demands of the system under study.
These approximation techniques enable researchers to predict aspects of chemical reactivity by estimating the net energy released or absorbed during a process and mapping the pathway connecting reactants to products. As chemical bonds rupture and assemble, the system must typically traverse an activation barrier whose magnitude determines the reaction velocity at a specified thermal state. Nevertheless, predicting the outcome of combining two reactive reagents remains computationally demanding, as simulations must account for competing reaction channels, solvent environments, temperature effects, and potential catalytic influences that single static calculations rarely resolve entirely.
The distinguishing characteristic of the vibe-qc framework lies in its automated generation through artificial intelligence agents operating as supervised programmers. Under Peintinger’s scientific direction, these digital agents drafted algorithms and assembled the modular architecture of the codebase, completing a comprehensive initial version within ten weeks.
Rigorous human validation proved indispensable throughout the automated coding process, as generative assistants frequently produced syntactically valid code containing fundamental physical errors. On the initial day of code generation, for instance, an assistant proposed modeling a neutral lithium atom using a computational method restricted to paired-electron configurations, overlooking the third unpaired valence electron characteristic of the element. While simple electron counting revealed that initial discrepancy, more nuanced programming defects were identified only by systematically benchmarking the output of vibe-qc routines against independently authored quantum chemistry packages.
The investigation also highlighted how computational predictions diverge depending on the level of physical approximation embedded within a simulation model. When evaluating the initiation energy required for chemical reactions, calculations derived from highly simplified models exhibited substantial average deviations when compared to values produced by complex electronic-structure methods. Because both sets of conflicting values represented synthetic computer outputs, Peintinger underscored that laboratory experimental measurements remain the definitive empirical standard for verifying the physical realism of any computational prediction.
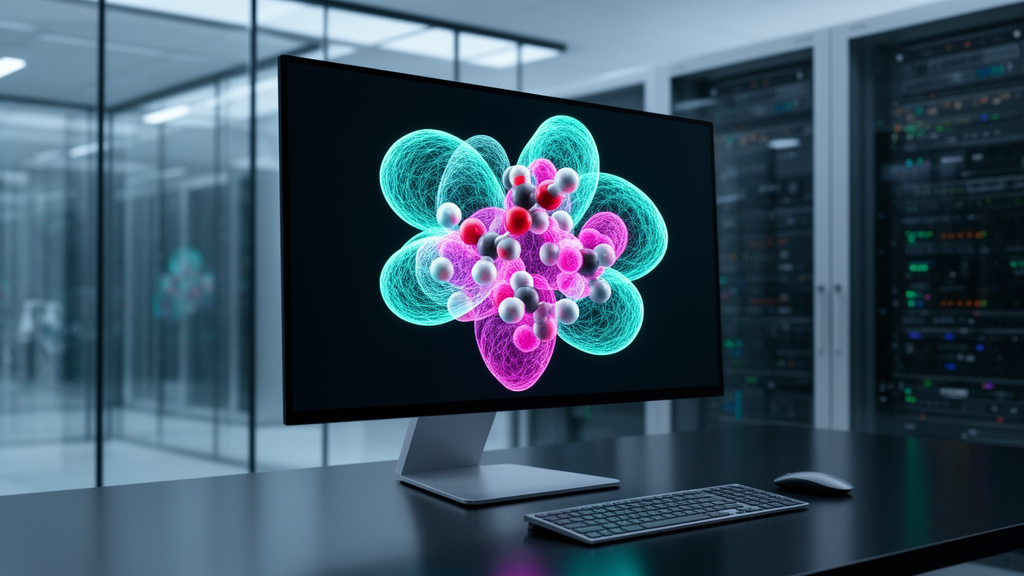
Currently, vibe-qc encompasses a versatile suite of computational methods capable of handling both discrete molecular structures and periodic crystalline materials, spanning computationally lightweight approximations to computationally demanding, high-precision algorithms. The entire software package and its associated tools, including the vibe-viewer visualization utility used to render electronic probability clouds and phase signs, are released as open-source software under the Mozilla Public License, granting the scientific community complete freedom to examine, adapt, and distribute the underlying codebase.
Peintinger structured vibe-qc around an automated development workflow where generative artificial intelligence agents functioned as digital programmers working under direct, continuous scientific oversight. Rather than creating entirely new mathematical formulas for electronic structure, the research focused on this novel operational workflow, orchestrating artificial intelligence agents to draft algorithms and execute unit tests rapidly. Within approximately ten weeks, the collaborative setup produced a comprehensive initial software framework capable of supporting computational modeling across diverse chemical systems. Over a period of several months, this foundation expanded into a general-purpose electronic-structure platform suited for both isolated molecular configurations and extended periodic crystal lattices. Consequently, the project demonstrates that an individual researcher can now direct and maintain a software suite of substantial complexity rather than writing the entire codebase from scratch through manual programming.
The necessity of meticulous human verification remained central throughout the coding phase, because generative systems can generate syntactically convincing code that violates foundational physical principles. During the very first day of software generation, for example, the automated assistant attempted to model a neutral lithium atom using a computational procedure that processes all electrons in paired arrangements. Because a lithium atom possesses three electrons, exactly one electron remains unpaired, making the proposed computational strategy physically invalid. Although the algorithmic proposal appeared plausible on the surface, identifying the underlying mistake required straightforward electron counting. More intricate programming inconsistencies did not produce runtime errors and could only be identified when numerical calculations were benchmarked against outputs produced by independently authored quantum chemistry packages. Under Peintinger’s protocol, computational methods within vibe-qc were considered ready for deployment only after their outputs stood up to rigorous comparative cross-testing against established external software.
The most demanding debugging challenges arose when vibe-qc executed without producing any system error messages and generated outwardly convincing numerical values that nevertheless disagreed with alternative computational programs. In one comparative analysis, two distinct software platforms produced different numerical outputs because they handled the inner core electrons of the atoms under different physical models. One calculation excluded certain inner electrons while the other accounted for them, meaning that the two packages were not evaluating the identical pool of electrons. Although both numerical results were mathematically correct within their respective algorithmic implementations, each corresponded to distinct physical questions. Resolving such computational discrepancies required the researcher to reexamine the foundational physical assumptions and approximations embedded within each model rather than searching for standard coding defects.
Assessing the reliability of theoretical calculations also demands an explicit understanding of how model simplifications influence final predictions. When calculating the activation energy required to initiate a chemical transformation, results generated by simplified computational models exhibited substantial average deviations from values derived from more complex, computationally demanding methods. Because both sets of conflicting numbers represented synthetic estimations generated entirely on a computer, computational comparison alone could not establish empirical accuracy. Peintinger stressed that determining how closely any simulated prediction reflects nature requires direct comparison with measured empirical values obtained from laboratory experiments. Theoretical models provide estimates of energy released or absorbed and trace hypothetical reaction pathways, but real-world chemical outcomes, such as the behavior observed when mixing reagents, involve competing pathways, solvent interactions, and thermal conditions that computation cannot fully resolve without experimental validation.
The modeling of molecular adsorption on solid surfaces provides a practical illustration of how electronic-structure calculations clarify surface-mediated phenomena, such as catalytic reactions. In investigating how water molecules interact with a thin monolayer of boron nitride, a material composed of boron and nitrogen atoms, researchers compute total system energies across diverse spatial coordinates and molecular orientations. By comparing these calculated energies, scientists identify the structural configuration with the lowest energy, which represents the most energetically favorable arrangement and reveals how firmly the molecule adheres to the substrate. Furthermore, associated software tools such as vibe-viewer enable the visualization of electron probability regions calculated by vibe-qc, rendering spatial distribution clouds where opposing colors represent opposite computational signs. Distributed openly under the Mozilla Public License, the complete codebase provides accessible computational simulation tools spanning rapid approximations to intricate electronic methods for investigating molecular and crystalline properties.
Subject of Research: Development of open-source quantum chemistry software using generative artificial intelligence
Article Title: Electrons reveal what will happen in the test tube
Article References: Electrons reveal what will happen in the test tube. Original publication (Bibliographic details unavailable.)
Image Credits: AI Generated
DOI: Not provided
Keywords: quantum chemistry, vibe-qc, generative AI, electronic structure, University of Bonn, Mulliken Center for Theoretical Chemistry, computational chemistry, open-source software, Mozilla Public License, boron nitride, surface adsorption, catalytic reactions, diamond crystals, reaction pathways, energy barriers, AI agents
Cite Scienmag News
Bethany Barker. (November 10, 2026). Generative AI System Develops Open-Source Quantum Chemistry Software in Ten Weeks. Scienmag. https://scienmag.com/generative-ai-system-develops-open-source-quantum-chemistry-software-in-ten-weeks/
Bethany Barker. "Generative AI System Develops Open-Source Quantum Chemistry Software in Ten Weeks." Scienmag, 10 November 2026, https://scienmag.com/generative-ai-system-develops-open-source-quantum-chemistry-software-in-ten-weeks/. Accessed 11 November 2026.
Bethany Barker. "Generative AI System Develops Open-Source Quantum Chemistry Software in Ten Weeks." Scienmag. November 10, 2026. https://scienmag.com/generative-ai-system-develops-open-source-quantum-chemistry-software-in-ten-weeks/




